Managing Change Controls Between a CDMO and a Sponsor/MAH

It is crucial for a Marketing Authorization Holder (MAH) to review and approve changes made by a Contract Development and Manufacturing Organization (CDMO) for several important reasons:

Regulatory Compliance

The Market Authorization Holder (MAH) – or the sponsor for pre-commercial GMP manufacturing – bears the primary responsibility for ensuring compliance with the marketing authorization and regulatory requirements throughout the product’s lifecycle. By reviewing and approving CDMO changes, the MAH can:

  • Ensure changes align with the approved marketing authorization
  • Verify that any variations to the marketing authorization are properly submitted to regulatory authorities
  • Maintain oversight of post-approval change management as required by regulations

Before I go any further on the topic I want you to go and read my post Classification of Changes for GMP/GDP. This post will build on that discussion.

I think it is better for the CDMO to put a lot of thought into this, and the MAH (the client) to evaluate and adapt. For all but the big players, the volume is going to be on the CDMO’s side. But if you are the client and your CDMO hasn’t taken this into account to the appropriate degree, you need to ensure appropriate steps taken. As such the rest of this post will be written from the CDMO’s side, but the same principles apply to the MAH (and should be included in the audit program).

Remember we have three goals:

  • Fulfill our contractual responsibilities
  • Help the MAH maintain appropriate control as the product owner
  • Ensure alignment between both parties on change implementation

The critical requirement here is ensuring the right changes get to the right client so they can be filled the right way. Returning to basics, we are approaching changes as:

Now it’s easy to apply this to product. Create and/or receive the design space and the control space. Everything that falls into a non-established condition does not get reported to the client at time of execution. If it is “Do and Report” is is in the APQR. If it is “Do and Record” they can see it during the audit.
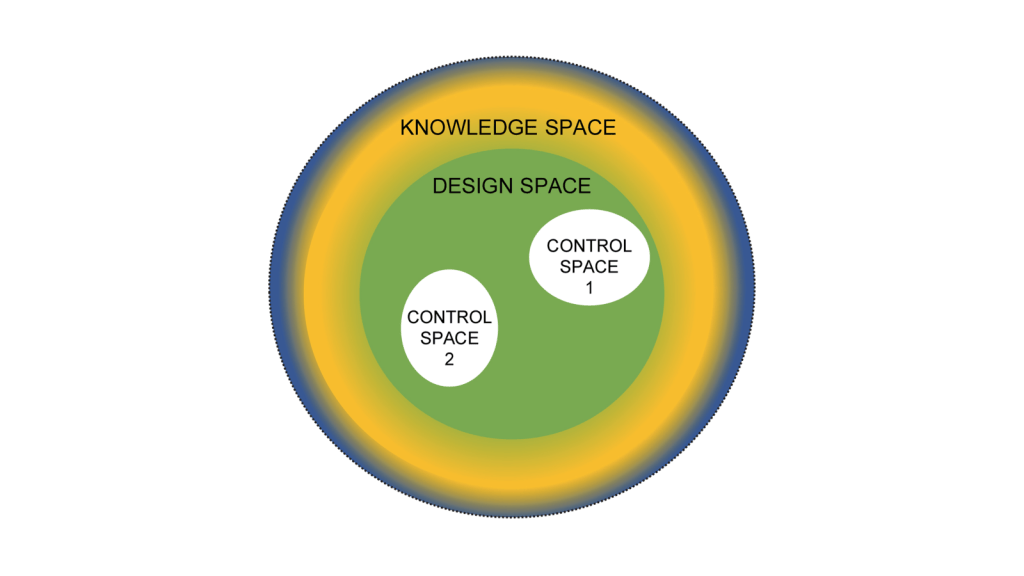
Where a lot of CDMOs trip up here are facility and quality system changes. My recommendation here is the same, define a design space based on the CMC section of the Common Technical Document which basically boils down to:

The CMC (Chemistry, Manufacturing, and Controls) section of a regulatory dossier typically includes the following key facility-related information:

  • Manufacturing Facilities
    • Names and addresses of all manufacturing, testing, and storage facilities involved in production
    • Description of the manufacturing operations performed at each site
    • Floor plans and layouts of production areas
    • Details on utilities and support systems (HVAC, water, gases, etc.)
    • Information on facility design features for contamination control and product protection
  • Equipment
    • List of major production and laboratory equipment
    • Equipment specifications and capacities
    • Cleaning and maintenance procedures for equipment
  • Environmental Controls
    • Description of clean room classifications and environmental monitoring programs
    • Air handling systems and controls
    • Water systems (purified water, water for injection) and controls
  • Material Flow
    • Personnel and material flow diagrams
    • Segregation of operations to prevent cross-contamination
  • Quality Control Laboratories
    • Description of QC lab facilities and equipment
    • Environmental controls in QC labs
  • Storage Areas
    • Description of storage facilities for raw materials, intermediates, and finished products
    • Storage conditions and controls (temperature, humidity, etc.)

There is a whole lot of wiggle room here in things that fall into “Do and Record.” By building this into your change control system you can delineate what goes to to the client and what doesn’t. I recommend sitting down with this list and deciding what types of changes fall into “Tell and Do” – what you ask permission from clients before doing; “Do and Report” – what goes in the APQR; and, “Do and Record” – what the client sees when they audit.

You know have good rules on what changes go to a client for prior approval and which ones do not. This gets codified in two places: the change control process and the quality/technical agreement.

Some other things to build into your change control process:

  1. Documenting when a client requests a change, the reason and the impact on the platform. Remember you have other clients, and more and more CDMO’s are offering a platform, so there needs to be appropriate review and endorsement.
  2. Think through how changes to facility (and other platform elements) are communicated and gated for multiple clients. Have a mechanism to manage client specific activities and to track first-product impacted for multiple products.
  3. Have clear timelines and expectations on change communication and approval with the client in the quality/technical agreement. Hold each other accountable.
  4. Have contingency plans. There will always be that one client who will be in shortage if you make that urgent change just when you want/need to.
  5. Have a method for evaluating requested changes to the change plan by clients and making decisions around it. There will be that one client who doesn’t agree or wants something weird that disagrees with what all the other clients want.
  6. Have rules in place to manage changes inactive for long periods or extensions specific for those changes that rise to client approval. These will have a different flow than internal changes.

I’ve used a bit of commercial headspace for this post, relying on the APQR. For clinical processes, product tends to fall into campaign-mindset, so “Do and Report” ends up being more a clinical campaign change report than an APQR.
Water, Water, Everywhere

XKCD, https://xkcd.com/2982/

Everyone probably feels like the above illustration sooner or later about their water system.

The Critical Role of Water in Pharmaceutical Manufacturing

In the pharmaceutical industry, we often joke that we’re primarily water companies that happen to make drugs on the side. This quip underscores a fundamental truth: water is a crucial component in drug manufacturing processes. Its purity and quality are paramount to ensuring the safety and efficacy of pharmaceutical products.

Why Water Quality Matters

Water is ubiquitous in pharmaceutical manufacturing, used in everything from cleaning equipment to serving as a key ingredient in many formulations. Given its importance, regulatory bodies like the FDA and EMA have established stringent Good Manufacturing Practice (GMP) guidelines for water systems in pharmaceutical facilities.

GMP Requirements for Water Systems

The GMPs mandate that water systems be meticulously designed, constructed, installed, commissioned, qualified, monitored, and maintained. The primary goal? Preventing microbiological contamination. This comprehensive approach encompasses several key areas:

  1. System Design: Water systems must be engineered to minimize the risk of contamination.
  2. Construction and Installation: Materials and methods used must meet high standards to ensure system integrity.
  3. Commissioning and Qualification: Rigorous testing is required to verify that the system performs as intended.
  4. Monitoring: Ongoing surveillance is necessary to detect any deviations from established parameters.
  5. Maintenance: Regular upkeep is crucial to maintain system performance and prevent degradation.

Key Regulatory Requirements

AgencyTitleYearURL
EMAGuideline on the quality of water for pharmaceutical use2020https://www.ema.europa.eu/en/documents/scientific-guideline/guideline-quality-water-pharmaceutical-use_en.pdf
WHOGood manufacturing practices: water for pharmaceutical use2012https://www.who.int/docs/default-source/medicines/norms-and-standards/guidelines/production/trs970-annex2-gmp-wate-pharmaceutical-use.pdf
US FDAGuide to inspections of high purity water systems2016https://www.fda.gov/media/75927/download
PIC/SInspection of utilities2014https://picscheme.org/docview/1941
US FDAWater for pharmaceutical use2014https://www.fda.gov/media/88905/download
USP<1231> Water for pharmaceutical purposes2020Not publicly available
USP<543> Water Conductivity2020Not publicly available
USP<85> Bacterial Endotoxins Test2020Not publicly available
USP<643> Total Organic Carbon2020Not publicly available
Ph. Eur.Monograph 0168 (Water for injections)2020Not publicly available
Ph. Eur.Monograph 0008 (Purified water)2020Not publicly available

Specific Measures for Contamination Prevention

To meet these GMP requirements, pharmaceutical manufacturers must implement several specific measures:

Minimizing Particulates

Particulate matter in water can compromise product quality and potentially harm patients. Filtration systems and regular cleaning protocols are essential to keep particulate levels in check.

Controlling Microbial Contamination

Microorganisms can proliferate rapidly in water systems if left unchecked. Strategies to prevent this include:

  • Regular sanitization procedures
  • Maintaining appropriate water temperatures
  • Implementing effective water treatment technologies (e.g., UV light, ozonation)

Preventing Endotoxin Formation

Endotoxins, produced by certain bacteria, can be particularly problematic in pharmaceutical water systems. Measures to prevent endotoxin formation include:

  • Minimizing areas where water can stagnate
  • Ensuring complete drainage of pipes
  • Regular system flushing

The Ongoing Challenge

Maintaining water quality in pharmaceutical manufacturing is not a one-time effort but an ongoing process. It requires constant vigilance, regular testing, and a commitment to continuous improvement. As regulations evolve and our understanding of potential contaminants grows, so too must our approaches to water system management.

Types of Water

These water types are defined and regulated by pharmacopeias such as the United States Pharmacopeia (USP), European Pharmacopoeia (Ph. Eur.), and other regional standards. Pharmaceutical manufacturers must adhere to the specific requirements outlined in these references to ensure water quality and safety in drug production.

Potable Water

Potable water, also known as drinking water, may be used for some pharmaceuticals bt is more commonly used in cosmetics. It can also be used for cleanings walls and floors in non-asceptic areas.

Key points:

  • Must comply with EPA standards or comparable regulations in the EU/Japan
  • Can be used to manufacture drug substances (bulk drugs)
  • Not suitable for preparing USP dosage forms or laboratory reagents

Purified Water (PW)

Purified water is widely used in pharmaceutical manufacturing for non-sterile preparations.

Specifications (USP <1231>):

  • Conductivity: ≤1.3 μS/cm at 25°C
  • Total organic carbon (TOC): ≤500 ppb
  • Microbial limits: ≤100 CFU/mL

Applications:

  • Non-parenteral preparations
  • Cleaning equipment for non-parenteral products
  • Preparation of some bulk chemicals

Water for Injection (WFI)

Water for Injection is used for parenteral drug products and has stricter quality standards.

Specifications (USP <1231>):

  • Conductivity: ≤1.3 μS/cm at 25°C
  • TOC: ≤500 ppb
  • Bacterial endotoxins: <0.25 EU/mL
  • Microbial limits: ≤10 CFU/100 mL

Production methods:

  • Distillation
  • Reverse osmosis (allowed by Ph. Eur. since 2017)

Sterile Water for Injection (SWFI)

SWFI is WFI that has been sterilized for direct administration.

Characteristics:

  • Sterile
  • Non-pyrogenic
  • Packaged in single-dose containers

Highly Purified Water (HPW)

Previously included in the European Pharmacopoeia, but now discontinued.

Type of WaterDescriptionUSP ReferenceEP Reference
Potable WaterMeets drinking water standards, used for early stages of manufacturingNot applicableNot applicable
Purified Water (PW)Used for non-sterile preparations, cleaning equipmentUSP <1231>Ph. Eur. 0008
Water for Injection (WFI)Used for parenteral products, higher purity than PWUSP <1231>Ph. Eur. 0169
Sterile Water for Injection (SWFI)WFI that has been sterilized for direct administrationUSP <1231>Ph. Eur. 0169
Bacteriostatic Water for InjectionContains bacteriostatic agents, for multiple-dose useUSP <1231>Ph. Eur. 0169
Sterile Water for IrrigationPackaged in single-dose containers larger than 1LUSP <1231>Ph. Eur. 1116
Sterile Water for InhalationFor use in inhalators, less stringent endotoxin levelsUSP <1231>Ph. Eur. 1116
Water for HemodialysisSpecially treated for use in hemodialysis, produced on-siteUSP <1231>Not specified

Additional relevant USP chapters:

  • USP <645>: Water for Pharmaceutical Purposes – Microbial Attributes
  • USP <85>: Bacterial Endotoxins Test

Always refer to the most current versions of the pharmacopoeial monographs and regulatory guidelines for detailed information.

Good Water System Design

Hygienic and Sanitary Design

The cornerstone of any good water system is its hygienic and sanitary design. This principle encompasses several aspects:

  • Smooth, cleanable surfaces: All surfaces in contact with water should be smooth, non-porous, and easily cleanable to prevent biofilm formation.
  • Self-draining components: Pipes and tanks should be designed to drain completely, eliminating standing water that could harbor microorganisms.
  • Accessibility: All parts of the system should be easily accessible for inspection, cleaning, and maintenance.

Material Selection

Choosing the right materials is crucial for maintaining water quality and system integrity:

  • Corrosion resistance: Use materials that resist corrosion, such as stainless steel (316L grade for high-purity applications) or appropriate food-grade plastics.
  • Smooth internal finish: Crevices are places where corrosion happens, electropolishing improves the resistance of stainless steel to corrosion.
  • Leachate prevention: Select materials that do not leach harmful substances into the water, even under prolonged contact or elevated temperatures.
  • Non-adsorptive surfaces: Avoid materials that may adsorb contaminants, which could later be released back into the water.

Microbial Control

Preventing microbial growth is essential for water system safety:

  • Elimination of dead legs: Design piping to avoid areas where water can stagnate and microorganisms can proliferate.
  • Temperature control: Maintain temperatures outside the optimal range for microbial growth (typically below 20°C or above 50°C).
  • Regular sanitization: Incorporate features that allow for effective and frequent sanitization of the entire system.

System Integrity

Ensuring the system remains sealed and leak-free is critical:

  • Proper sealing: Use appropriate gaskets and seals compatible with the system’s operating conditions.
  • Pressure testing: Implement regular pressure tests to identify and address potential leaks promptly.
  • Quality connections: Utilize sanitary fittings and connections designed for hygienic applications.

Cleaning and Sanitization Compatibility

The system must withstand regular cleaning and sanitization:

  • Chemical resistance: Choose materials and components that can tolerate cleaning and sanitizing agents without degradation.
  • Thermal stability: Ensure all parts can withstand thermal sanitization processes if applicable.
  • CIP/SIP design: Incorporate Clean-in-Place (CIP) or Steam-in-Place (SIP) features for efficient and thorough cleaning.

Capacity and Performance

Meeting output requirements while maintaining quality is crucial:

  • Proper sizing: Design the system to meet peak demand without compromising water quality or flow rates.
  • Redundancy: Consider incorporating redundant components for critical parts to ensure continuous operation.
  • Efficiency: Optimize the system layout to minimize pressure drops and energy consumption.

Monitoring and Control

Implement robust monitoring systems to ensure water quality:

  • Sampling points: Strategically place sampling ports throughout the system for regular quality checks.
  • Instrumentation: Install appropriate instruments to monitor critical parameters such as flow rate, pressure, temperature, and conductivity.
  • Control systems: Implement automated control systems to maintain consistent water quality and system performance.

Regulatory Compliance

Ensure the system design meets all relevant regulatory requirements:

  • Material compliance: Use only materials approved for contact with water in your specific application.
  • Documentation: Maintain detailed documentation of system design, materials, and operating procedures.
  • Validation: Conduct thorough system qualification to demonstrate consistent performance and quality.

By adhering to these principles, you can design a water system that not only meets your capacity requirements but also ensures the highest standards of safety and quality. Remember, good water system design is an ongoing process that requires regular review and updates to maintain its effectiveness over time.

The Challenge of Cleanroom Classification Harmonization

In the world of pharmaceutical manufacturing, cleanroom classifications play a crucial role in ensuring product quality and patient safety. However, a significant hurdle in the global harmonization of regulations has been a pain in our sides for a long time, that highlights the persistent differences between major regulatory bodies, including the FDA, EMA, and others, despite efforts to align through organizations like the World Health Organization (WHO) and the Pharmaceutical Inspection Co-operation Scheme (PIC/S).

The Current Landscape

United States Approach

In the United States, cleanroom classifications are primarily governed by two key documents:

  1. The FDA’s “Sterile Drug Products Produced by Aseptic Processing” guidance
  2. ISO 14644-1 standard for cleanroom classifications

The ISO 14644-1 standard is particularly noteworthy as it’s a general standard applicable across various industries utilizing cleanrooms, not just pharmaceuticals.

European Union Approach

The European Union takes a different stance, employing a grading system outlined in the EU GMP guide:

  • Grades A through D are used for normal cleanroom operation
  • ISO 14644 is still utilized, but primarily for validation purposes

World Health Organization Alignment

The World Health Organization (WHO) aligns with the European approach, adopting the same A to D grading system in its GMP guidelines.

The Implications of Disharmony

This lack of harmonization in cleanroom classifications presents several challenges:

  1. Regulatory Complexity: Companies operating globally must navigate different classification systems, potentially leading to confusion and increased compliance costs.
  2. Technology Transfer Issues: Transferring manufacturing processes between regions becomes more complicated when cleanroom requirements differ.
  3. Inspection Inconsistencies: Differences in classification systems can lead to varying interpretations during inspections by different regulatory bodies.

The Missed Opportunity in Annex 1

The recent update to Annex 1, a key document in GMP regulations, could have been a prime opportunity to address this disharmony. However, despite involvement from WHO and PIC/S (and through them the FDA), the update failed to bring about the hoped-for alignment in cleanroom classifications.

Moving Forward

As the pharmaceutical industry continues to globalize, the need for harmonized regulations continues to be central. I would love to see future efforts towards harmonization here that would:

  1. Prioritize alignment on fundamental technical specifications like cleanroom classifications
  2. Consider the practical implications for manufacturers operating across multiple jurisdictions

While the journey towards full regulatory harmonization may be long and challenging, addressing key discrepancies like cleanroom classifications would represent a significant step forward for the global pharmaceutical industry.

FDA Nitrosamine Impurities Update

FDA guidance, “Control of Nitrosamine Impurities in Human Drugs,” revises the final guidance of the same name issued on February 24, 2021, by including information about nitrosamine drug substance related impurities (NDSRIs), recommending implementation of new nitrosamine control strategies, and providing an updated timeline for manufacturers and applicants to implement these recommendations.

Nitrosamine impurities are important to control because they are potential human carcinogens. Long-term exposure to these impurities at levels above acceptable limits can increase the risk of cancer. Nitrosamines can be found in various consumer products and the environment, and they have been detected in several pharmaceutical products since 2018, prompting recalls and regulatory actions. A lot of regulatory action. Nitrosamine impurities may be one of the biggest drivers of changes in the GMPs.

Current Regulatory View

Regulators, including the FDA, Health Canada, and the European Medicines Agency (EMA), have been actively working to address the presence of nitrosamine impurities in medications. The current regulatory view emphasizes:

  • Risk Assessment and Control: Regulatory agencies have established acceptable intake (AI) limits for nitrosamines in drug products. These limits are designed to minimize the risk of cancer associated with long-term exposure to these impurities.
  • Guidance and Frameworks: Agencies have issued guidance documents outlining frameworks for assessing and controlling nitrosamine impurities. For example, the FDA’s guidance includes recommendations for predicting the mutagenic and carcinogenic potential of nitrosamine drug substance-related impurities (NDSRIs) and provides AI limits based on carcinogenic potency categorization.
  • International Collaboration: There is significant collaboration among global regulators to harmonize approaches and methodologies for controlling nitrosamine impurities. This includes the adoption of the Carcinogenic Potency Categorization Approach (CPCA) to determine AI limits.
  • Industry Responsibility: Manufacturers are responsible for understanding their processes to prevent nitrosamine formation and for conducting risk assessments. They must implement control strategies and perform confirmatory testing to ensure that nitrosamine levels remain below the established AI limits.

Regulators are focused on ensuring the safety of pharmaceutical products by controlling nitrosamine impurities through comprehensive risk assessments, setting stringent AI limits, and fostering international cooperation. Companies need to make sure they are ahead of this matter.

Viral Controls in Facility Design

Facility design and control considerations for mitigating viral contamination risk is a holistic approach to facility design and controls, considering all potential routes of viral introduction and spread. A living risk management approach should be taken to identify vulnerabilities and implement appropriate mitigation measures.

Facility Considerations

  • Segregation of areas: Separate areas for cell banking, small-scale and large-scale upstream cell culture/fermentation, downstream processing, media/buffer preparation, materials management, corridors, and ancillary rooms (e.g. cold rooms, freezer rooms, storage areas).
  • Traffic flow: Control and minimize traffic flow of materials, personnel, equipment, and air within and between areas and corridors. Implement room segregation strategies.
  • Air handling systems: Design HVAC systems to maintain appropriate air quality and prevent cross-contamination between areas. Use HEPA filtration where needed.
  • Room Classifications
    • For open operations:
      • Open sterile and aseptic operations must be performed in an environment where the probability of contamination is acceptably low, i.e. an environment meeting the bioburden requirements for a Grade A space.
      • Open bioburden-controlled processing may be performed in an ISO Grade 8/EU Grade C or EU Grade D environment as appropriate for the unit operation.
      • Open aseptic operations require a Grade A environment. Maintaining a Grade A cleanroom for large bioreactors is not feasible.
    • For closed operations:
      • Closed systems do not require cleanroom environments. ICH Q7 states that closed or contained systems can be located outdoors if they provide adequate protection of the material.
      • When all equipment used to manufacture a product is closed, the surrounding environment becomes less critical. The cleanroom requirements should be based on a business risk assessment and could be categorized as unclassified.
      • Housing a closed aseptic process in a Grade C or Grade B cleanroom would not mitigate contamination risk compared to an unclassified environment.
      • For low bioburden closed operations, the manufacturing environment can be unclassified.

Equipment Considerations

Closed vs. open processing: Utilize closed processing operations where possible to prevent introduction/re-introduction of viruses. Implement additional controls for open processing steps.

Closure LevelDescription
Closed EquipmentSingle use, never been used, such as irradiated and autoclaved assembles; connections are made using sterile connectors or tube wielders/sealers
Functionally closed equipment: cleaned and sterilizedOpen vessels or connections that undergo cleaning and sterilization prior to use and are then aseptically connected. The connection is then sterilized after being closed and remains closed during use.
Functionally closed equipment: cleaned and sanitizedOpen vessels or connections that are CIPed including bioburden reducing flushes, but not sterilized before use and remain closed during use
OpenConnections open to the environment without subsequent cleaning, sanitization or sterilization prior to use

Operational Practices

  • Personnel controls: Implement rigorous training programs, safety policies and procedures for personnel working in critical areas.
  • Cleaning and sanitization: Establish frequent and thorough cleaning protocols for facilities, equipment, and processing areas using appropriate cleaning agents effective against viruses.
  • Material and equipment flow: Define procedures for disinfection and transfer of materials and equipment between areas to prevent contamination spread.
  • Storage practices: Implement proper storage procedures for product contact materials, intermediates, buffers, etc. Control access to cold rooms and freezers.

Additional Controls

  • Pest control: Implement comprehensive pest control strategies both inside and outside facilities, including regular treatments and monitoring.
  • Water systems: Design and maintain water systems to prevent microbial growth and contamination.
  • Process gases: Use appropriate filtration for process air and gases.
  • Environmental monitoring: Establish environmental monitoring programs to detect potential contamination early.