The State of the Analytical Lifecycle

There have been a lot of changes in the way pharma thinks of analytical lifecycles in the last few years. With changes in technology, new product modalities, ICH Q2(R2) and ICH Q14 being released in November 2023, and USP <1220> in 2022, it is fair to say we are all catching up with our analytical lifecycle programs.

Let’s discuss what I think are the four pivotal documents that provide direction.

ICH Q2(R2) and ICH Q14

ICH Q2(R2) and ICH Q14 are complementary guidelines that provide a comprehensive framework for the development, validation, and lifecycle management of analytical procedures used in the pharmaceutical industry.

ICH Q14 describes the scientific principles and risk-based approaches for developing and maintaining suitable analytical procedures throughout their lifecycle. It outlines the key elements and considerations for analytical procedure development, including:

  • Defining an Analytical Target Profile (ATP)
  • Knowledge management and risk assessment
  • Evaluating robustness and parameter ranges
  • Establishing an Analytical Procedure Control Strategy
  • Lifecycle management and post-approval changes
  • Multivariate analytical procedures
  • Real-time release testing

On the other hand, ICH Q2(R2) provides specific guidance on validating analytical procedures to demonstrate their suitability for the intended use. It covers various validation tests, methodologies, and evaluation criteria, such as:

  • Specificity/selectivity
  • Working range
  • Accuracy and precision
  • Robustness
  • Stability-indicating properties
  • Multivariate analytical procedures

In summary, ICH Q14 establishes the overarching principles and approaches for analytical procedure development. At the same time, ICH Q2(R2) focuses on the validation aspects to ensure the analytical procedures are fit for purpose and meet quality requirements throughout their lifecycle. The two guidelines are intended to be applied together, with ICH Q14 providing the framework for development and ICH Q2(R2) specifying the validation requirements.

USP <1220> Analytical Procedure Lifecycle and USP <1058> Analytical Instrument Qualification

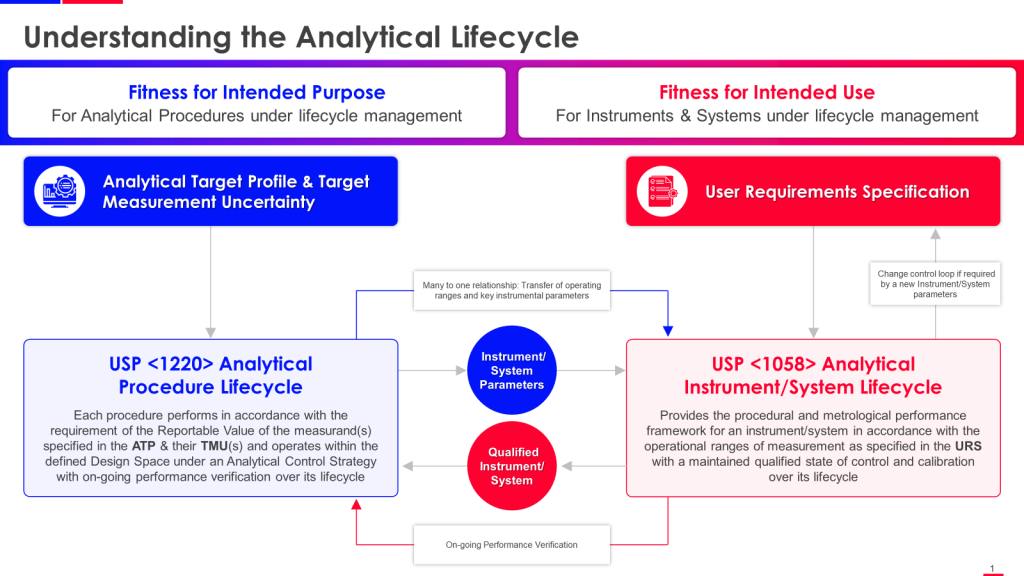
USP <1220> Analytical Procedure Lifecycle and USP <1058> Analytical Instrument Qualification are closely connected and complementary guidelines that provide a comprehensive framework for ensuring data integrity and quality in analytical procedures throughout their lifecycle.

The key connections between USP <1220> and USP <1058> are:

  1. USP <1220> establishes the principles and requirements for managing the entire lifecycle of analytical procedures, from procedure design and development to retirement. It emphasizes the importance of defining an Analytical Target Profile (ATP) and implementing an Analytical Procedure Control Strategy.
  2. USP <1058> focuses explicitly on the qualification of analytical instruments that execute analytical procedures. It outlines the requirements for ensuring instruments are suitable for their intended use through proper qualification (Design, Installation, Operational, and Performance Qualification).
  3. The instrument qualification activities described in USP <1058> are critical to the overall Analytical Procedure Control Strategy outlined in USP <1220>. Proper instrument qualification as per <1058> helps ensure the quality and integrity of data generated by analytical procedures throughout their lifecycle.
  4. Both guidelines stress the importance of defining user requirements (ATP in <1220> and User Requirements Specification in <1058>) as the basis for procedure development and instrument qualification activities.
  5. USP <1220> requires ongoing monitoring and periodic requalification of analytical procedures, which includes re-evaluating the suitability of the analytical instruments used, as described in the Performance Qualification section of <1058>.

USP <1220> provides the overarching framework for holistically managing analytical procedures. USP <1058> focuses on ensuring the instruments used to execute those procedures are properly qualified and suitable for their intended use. The two guidelines work together to maintain data integrity and quality across the entire analytical lifecycle.
Complementary Approaches

USP <1220> Analytical Procedure Lifecycle is closely related to and complements the ICH Q2(R2) and ICH Q14 guidelines.

  1. USP <1220> aligns with the principles outlined in ICH Q14 for managing the entire lifecycle of analytical procedures, from design and development to retirement. Both emphasize defining an Analytical Target Profile and implementing an Analytical Procedure Control Strategy.
  2. The validation activities described in ICH Q2(R2), such as evaluating specificity, accuracy, precision, and robustness, are critical components of the Analytical Procedure Control Strategy required by USP <1220>.
  3. USP <1220> requires ongoing monitoring and periodic requalification of analytical procedures, which aligns with the lifecycle management approach promoted in ICH Q14 and the validation during the lifecycle section in Q2(R2).
  4. All these guidelines stress the importance of knowledge management, risk management, and a science/risk-based approach throughout the analytical procedure lifecycle.
  5. The instrument qualification requirements outlined in USP <1058> are an integral part of the overall Analytical Procedure Control Strategy described in USP <1220>, ensuring instruments are suitable as per ICH Q2(R2) validation principles.

In essence, USP <1220> provides a comprehensive framework for analytical procedure lifecycle management that incorporates and operationalizes the scientific principles and validation activities detailed in the ICH Q14 and Q2(R2) guidelines, while USP <1058> provides the roadmap for instrument qualification. These four documents establish harmonized best practices for analytical procedures from development through retirement.
Leveraging Inspection Manuals for GMP Inspection Readiness

The various agency inspection manuals are critical tools for inspection readiness. I want to lay out where to find some of these manuals and then go deep into pre-approval inspections, focusing on data integrity.

European Medicines Agency

The European Medicines Agency (EMA) has established detailed procedures and work instructions for coordinating and conducting Good Clinical Practice (GCP), Good Manufacturing Practice (GMP), and pharmacovigilance inspections. Here are the key points regarding EMA’s inspection procedures:

GCP Inspection Procedures

  • EMA identifies applications for GCP inspections based on risk assessment criteria and exchanges information on shared applications with the FDA.
  • Inspections can be joint (conducted concurrently by EMA and FDA inspectors) or sequential (conducted separately by each agency).
  • EMA notifies the applicant/marketing authorization holder (MAH) and inspects sites about upcoming inspections through the IRIS industry portal instead of formal letters.
  • Applicants/MAHs must provide a signed statement accepting the inspection and granting direct access to documents and medical records.
  • Requested documents should be provided directly to inspectors in electronic format after consulting the reporting inspector.
  • After the inspection, EMA receives the draft inspection report, finalizes it with the inspectee’s responses, and publishes it in IRIS.

GMP Inspection Procedures

  • EMA coordinates GMP inspections based on risk assessment for marketing authorization applications, variations, and routine re-inspections.
  • Work instructions cover areas such as inspection announcement, fee calculation, product sampling/testing, and report circulation.

Pharmacovigilance Inspection Procedures

  • EMA has specific procedures for coordinating pharmacovigilance inspections and managing non-compliance notifications from MAHs.
  • Work instructions detail the inspection program creation, data entry in databases, and interactions with third-country inspectorates.

The EMA aims to harmonize inspection processes with the FDA and other regulatory bodies to streamline collaboration and information sharing while ensuring clinical trial subject protection and product quality.

FDA

The FDA Investigations Operations Manual (IOM) is the primary inspection manual used by FDA personnel when performing inspections and investigations.

The key points about the IOM are:

  • It provides comprehensive instructions, procedures, and policies for FDA investigators and inspectors to follow when conducting inspections, surveys, and investigations.
  • It covers inspectional activities for foods, drugs, medical devices, biologics, cosmetics, and other FDA-regulated products.
  • The manual details procedures for inspections of manufacturing facilities, sampling, import operations, recalls, consumer complaints, and other compliance activities.
  • It aims to ensure inspections are conducted consistently across FDA field offices and provide clear guidance to the industry on the FDA’s inspection approach.
  • The IOM is updated periodically to incorporate new laws, regulations, policies, and technological changes impacting FDA’s operations.
  • While not legally binding, the IOM represents the FDA’s current thinking and policies on inspections and investigations.

The FDA Investigations Operations Manual serves as the comprehensive inspection reference and procedure manual for FDA field staff carrying out the agency’s oversight and enforcement activities across all regulated product areas.

Pre-Approval Inspections

For new facilities, CPGM 7346.832, the FDA’s Compliance Program Guidance Manual for Pre-Approval Inspections (PAIs) of drug manufacturing facilities, is critical to spend time with. It outlines the objectives and procedures for FDA inspectors to evaluate a facility’s readiness for commercial manufacturing before approving a new drug application.

The key objectives of CPGM 7346.832 are:

  1. Assess if the facility has a quality system capable of controlling commercial manufacturing operations.
  2. Verify that the manufacturing processes, formulation, and analytical methods conform to the application details.
  3. Audit raw data integrity to authenticate the data submitted in the application.
  4. Evaluate the facility’s commitment to quality in pharmaceutical development (new objective added in 2022 revision).

The guidance instructs inspectors on evaluating the firm’s quality systems, process validation, data integrity, laboratory controls, change management, investigations, batch release procedures, and compliance with current Good Manufacturing Practices (cGMPs). It aims to ensure the facility can reliably produce the drug product described in the application.

Data Integrity

CPGM 7346.832 has specific requirements for data integrity audits during drug manufacturing facility pre-approval inspections (PAIs). Utilizing this document is an excellent way to evaluate your data integrity program.

The key points are:

  1. Objective 3 of the guidance is “Data Integrity Audit”—auditing and verifying raw data associated with the product to authenticate the data submitted in the application.
  2. Inspectors must audit the accuracy and completeness of data reported by the facility for the product. This involves verifying the factual integrity (data matches what was submitted) and contextual integrity (supporting data is complete).
  3. Inspectors should examine raw data, such as chromatograms, analyst notebooks, electronic data, etc., and compare it to the summary data in the application’s Chemistry, Manufacturing, and Controls (CMC) section.
  4. The data integrity audit should focus on finished product stability, dissolution, content uniformity, API impurities, etc.
  5. Inspectors must identify any unreported relevant data, data falsification, improper invalidation of results, or unexplained data discrepancies.
  6. Indications of data integrity issues include altered raw data, references to failing studies, discrepancies between samples, and missing records.

The data integrity audit aims to ensure the CMC data submitted to FDA is complete, reliable, and can be fully authenticated from the raw data at the manufacturing site. Robust data integrity is critical for the FDA to decide on the application’s approval.

Commissioning, Qualification and Validation

Commissioning, qualification, and validation are three distinct but interrelated processes in the pharmaceutical and biotechnology industries that ensure facilities, equipment, systems, and processes meet regulatory requirements and produce products of the desired quality. Here are the key differences:

Commissioning

  • Commissioning is a systematic process of ensuring that equipment, systems, and facilities are designed, installed, and functioning according to operational and engineering requirements.
  • It involves design reviews, installation verification, functional testing, and handover to operations.
  • Commissioning primarily focuses on satisfying engineering requirements and does not have direct regulatory requirements.

Qualification

  • Qualification is a regulated and documented process that demonstrates that equipment, systems, and facilities are installed correctly and operate as intended for their specific use.
  • It applies only to equipment, systems, and utilities that directly or indirectly impact product quality and patient safety.
  • Qualification activities include Installation Qualification (IQ), Operational Qualification (OQ), and Performance Qualification (PQ).
  • Qualification is focused on by regulatory authorities like the FDA and EMA to ensure compliance.

Validation

  • Validation is a broader concept establishing documented evidence that a process consistently produces a product that meets its predetermined specifications and quality attributes.
  • It encompasses the entire process lifecycle, including process design, qualification of equipment/systems, and continued process verification.
  • Validation ensures that the equipment and systems are qualified and the entire process is controlled to produce the desired final product.

In summary, commissioning verifies engineering requirements, qualification demonstrates suitability for intended use, and validation provides a high degree of assurance that the process will consistently produce a quality product. These activities are interconnected, with commissioning often leveraged during qualification and qualification being a subset of the overall validation process.

FDA’s Framework for Process Validation

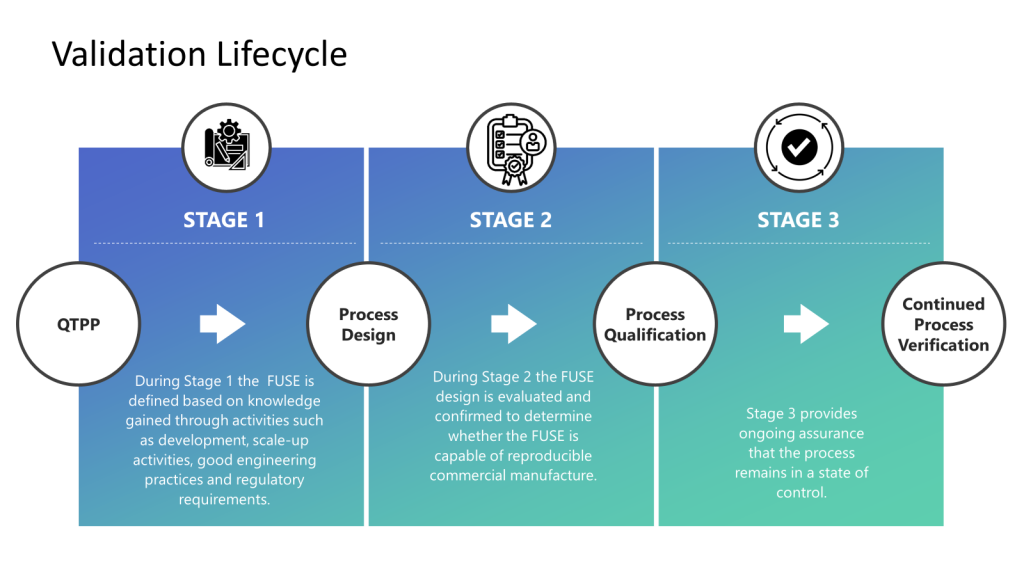
The FDA’s Process Validation Guidance is a core document outlining a lifecycle approach with outlines a lifecycle approach with three main stages:

Stage 1: Process Design

  • Establish a process design based on knowledge gained through development and scale-up activities.
  • Identify critical quality attributes (CQAs) and critical process parameters (CPPs) using risk assessment and multivariate studies like Design of Experiments (DoE).
  • Develop a control strategy to ensure CQAs are met.

Stage 2: Process Qualification

  • Evaluate the process design through facility, utility, and equipment qualification.
  • Conduct performance qualification (PQ) by running production batches to confirm the process design has reproducible commercial manufacturing.
  • Establish scientific evidence that the process meets all defined requirements and product specifications.

Stage 3: Continued Process Verification

  • Maintain the validated status and monitor performance to ensure a state of control.
  • Identify sources of variation and implement process improvements through an ongoing program.
  • Conduct product quality reviews periodically to evaluate process performance.

The guidance emphasizes using a science and risk-based approach throughout the lifecycle, leveraging process understanding and knowledge gained from development through commercial production. Effective process validation requires good planning, documented evidence, and a robust quality system.

Crafting Good User Requirements

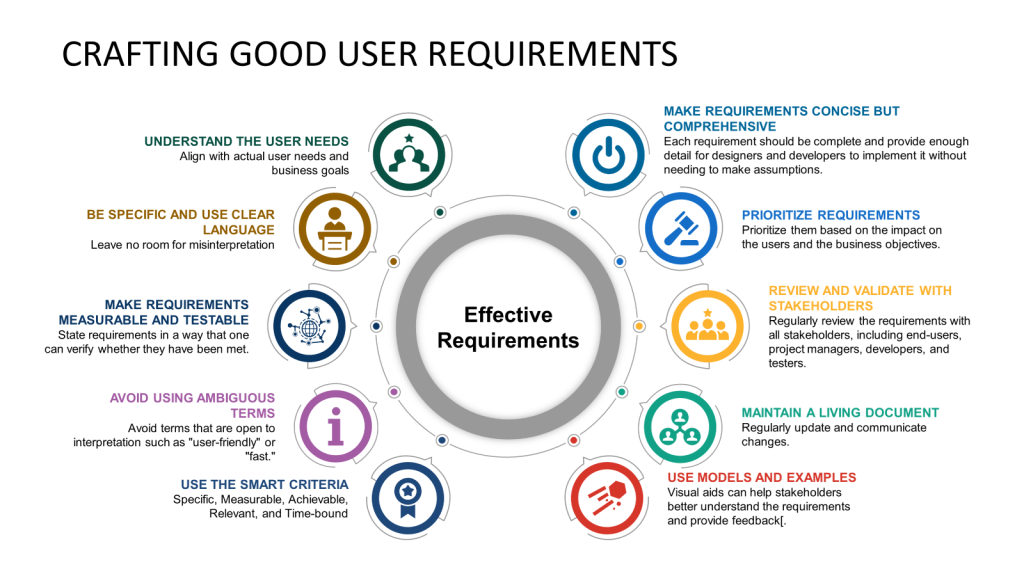
The User Requirements are a foundational document identifying the system’s product and process requirements. These product quality-related user requirements are based on product knowledge (CQAs), process knowledge (CPPs), regulatory requirements, and organization/site quality requirements. Writing a good user requirement for quality requirements involves several critical steps to ensure clarity, specificity, and effectiveness.

Understand the User Needs

Start by thoroughly understanding the user’s needs. This involves engaging with the end users or stakeholders to gather insights about their expectations, pain points, and the context in which the system will be used. This foundational step ensures that the requirements you develop are aligned with actual user needs and business goals.

Be Specific and Use Clear Language

Requirements should be specific and clearly stated to avoid ambiguity. Use simple, direct language and avoid technical jargon unless it is widely understood by all stakeholders. Define all terms and ensure that each requirement is phrased in a way that leaves no room for misinterpretation.

Make Requirements Measurable and Testable

Each requirement should be measurable and testable. This means stating requirements so one can verify whether they have been met. For example, instead of saying, “The system should load fast,” specify, “The system should load within 3 seconds when the number of simultaneous users is less than 10,000”.

Avoid Using Ambiguous Terms

Avoid terms open to interpretation, such as “user-friendly” or “fast.” If such terms are necessary, clearly define what they specifically mean in the context of your project. For instance, “user-friendly” is “the user can complete the desired task with no more than three clicks”.

Use the SMART Criteria

Employ the SMART criteria to ensure that each requirement is Specific, Measurable, Achievable, Relevant, and Time-bound. This approach helps set clear expectations and facilitates easier validation and verification of the requirements.

Make Requirements Concise but Comprehensive

While keeping each requirement concise and to the point is important, ensure all necessary details are included. Each requirement should be complete and provide enough detail for designers and developers to implement without making assumptions.

Prioritize Requirements

Not all requirements are equally important. Prioritize them based on their impact on the users and the business objectives. This helps manage the project scope and focuses on delivering maximum value.

It is good to categorize the user requirements here, such as:

  • Quality
  • Business
  • Health, Safety, and Environmental (HSE)

Review and Validate with Stakeholders

Review the requirements regularly with all stakeholders, including end-users, project managers, developers, and testers. This collaborative approach helps identify gaps or misunderstandings early in the project lifecycle.

Maintain a Living Document

Requirements might evolve as new information emerges or business needs change. Maintain your requirements document as a living document, regularly update it, and communicate changes to all stakeholders.

Use Models and Examples

Where applicable, use diagrams, mock-ups, or prototypes to complement the written requirements. Visual aids can help stakeholders better understand the requirements and provide feedback.

When writing user requirements for quality requirements, it’s crucial to avoid common pitfalls that can lead to misunderstandings, scope creep, and, ultimately, a product that does not meet the needs of the users or stakeholders. Here are some of the most common mistakes to avoid:

Ambiguity and Lack of Clarity

One of the most frequent errors in writing requirements is ambiguity. Requirements should be clear and concise, with no room for interpretation. Using vague terms like “user-friendly” or “fast” without specific definitions can lead to unmet expectations because different people may interpret these terms differently.

Incomplete Requirements

Another common issue is incomplete requirements that do not capture all necessary details or scenarios. This can result in features that do not fully address the users’ needs or require costly revisions later in development.

Overlooking Non-Functional Requirements

Focusing solely on what the system should do (functional requirements) without considering how it should perform (non-functional requirements), such as performance, security, and usability, can jeopardize the system’s effectiveness and user satisfaction.

Failure to Involve Stakeholders

Not involving all relevant stakeholders in the requirements gathering and validation process can lead to missing critical insights or requirements important to different user groups. This often results in a product that does not fully meet the needs of all its users.

Scope Creep

Without a clear definition of scope, projects can suffer from scope creep, where additional features and requirements are added without proper review, leading to delays and budget overruns. It’s important to have a well-defined project scope and a change management process in place.

Not Prioritizing Requirements

Not all requirements are equally important. Failing to prioritize requirements can misallocate resources and efforts on less critical features. Using prioritization techniques like MoSCoW (Must have, Should have, Could have, Won’t have this time) can help manage and focus efforts on what truly matters.

Lack of Validation and Verification

Skipping the validation (ensuring the product meets the intended use and needs of the stakeholders) and verification (ensuring the product meets the specified requirements) processes can lead to a final product not aligned with user needs and expectations.

Poor Documentation and Traceability

Inadequate documentation and lack of traceability can lead to confusion and errors during development. Maintaining detailed documentation and traceability from requirements through to implementation is crucial to ensure consistency and completeness.

Ignoring the Importance of Clear Communication

Effective communication is essential throughout the requirements process. Miscommunication can lead to incorrect or misunderstood requirements being developed. Regular, clear communication and documentation updates are necessary to keep all stakeholders aligned.

 Not Considering the Testing of Requirements

Considering how requirements will be tested during the definition phase is important. This consideration helps ensure that requirements are testable and that the final product will meet them. Planning for testing early can also highlight any potential issues with clarity or feasibility of requirements.

Validation Planning – A visual approach

Normally, when I write a blog post, I include the graphics, but I decided to separate them out to show some of my thought processes for designing slides.

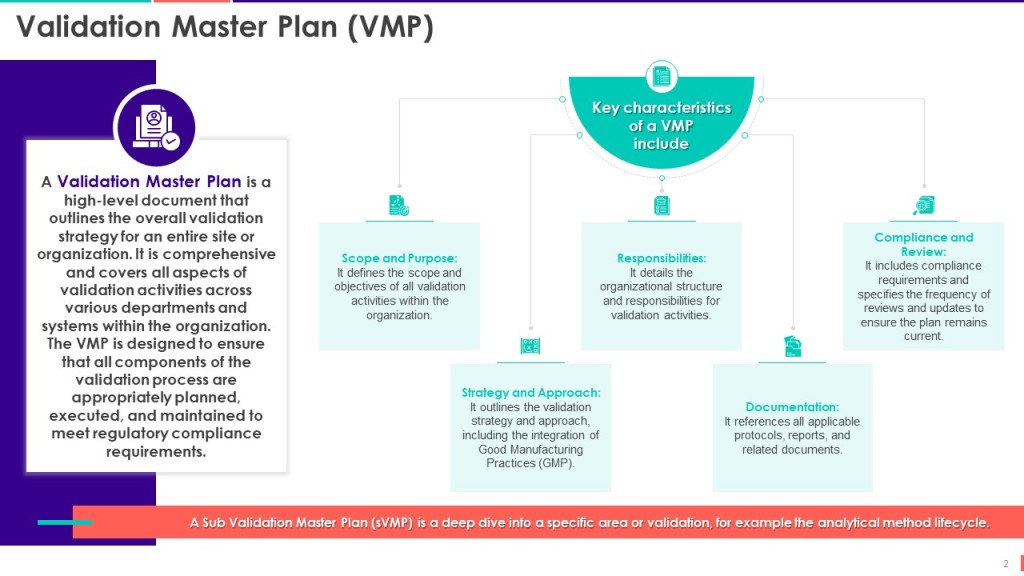
I start with a nice slide that introduces the topic I am going to discuss, introducing the main concept, the Validation Master Plan (VMP), and the Validation Plan (VP.)

My next slide details the Validation Master Plan in more depth, covering the VMP’s core characteristics.

Then dive into the reasons for having a VMP.

Then cover the Validation plan characteristics.

These are still rather wordy, and I think the last slide can be divided into two. But I have a pretty good training here.